We build the record that holds up.
Quality & regulatory compliance for pharmaceutical and medical device manufacturers.
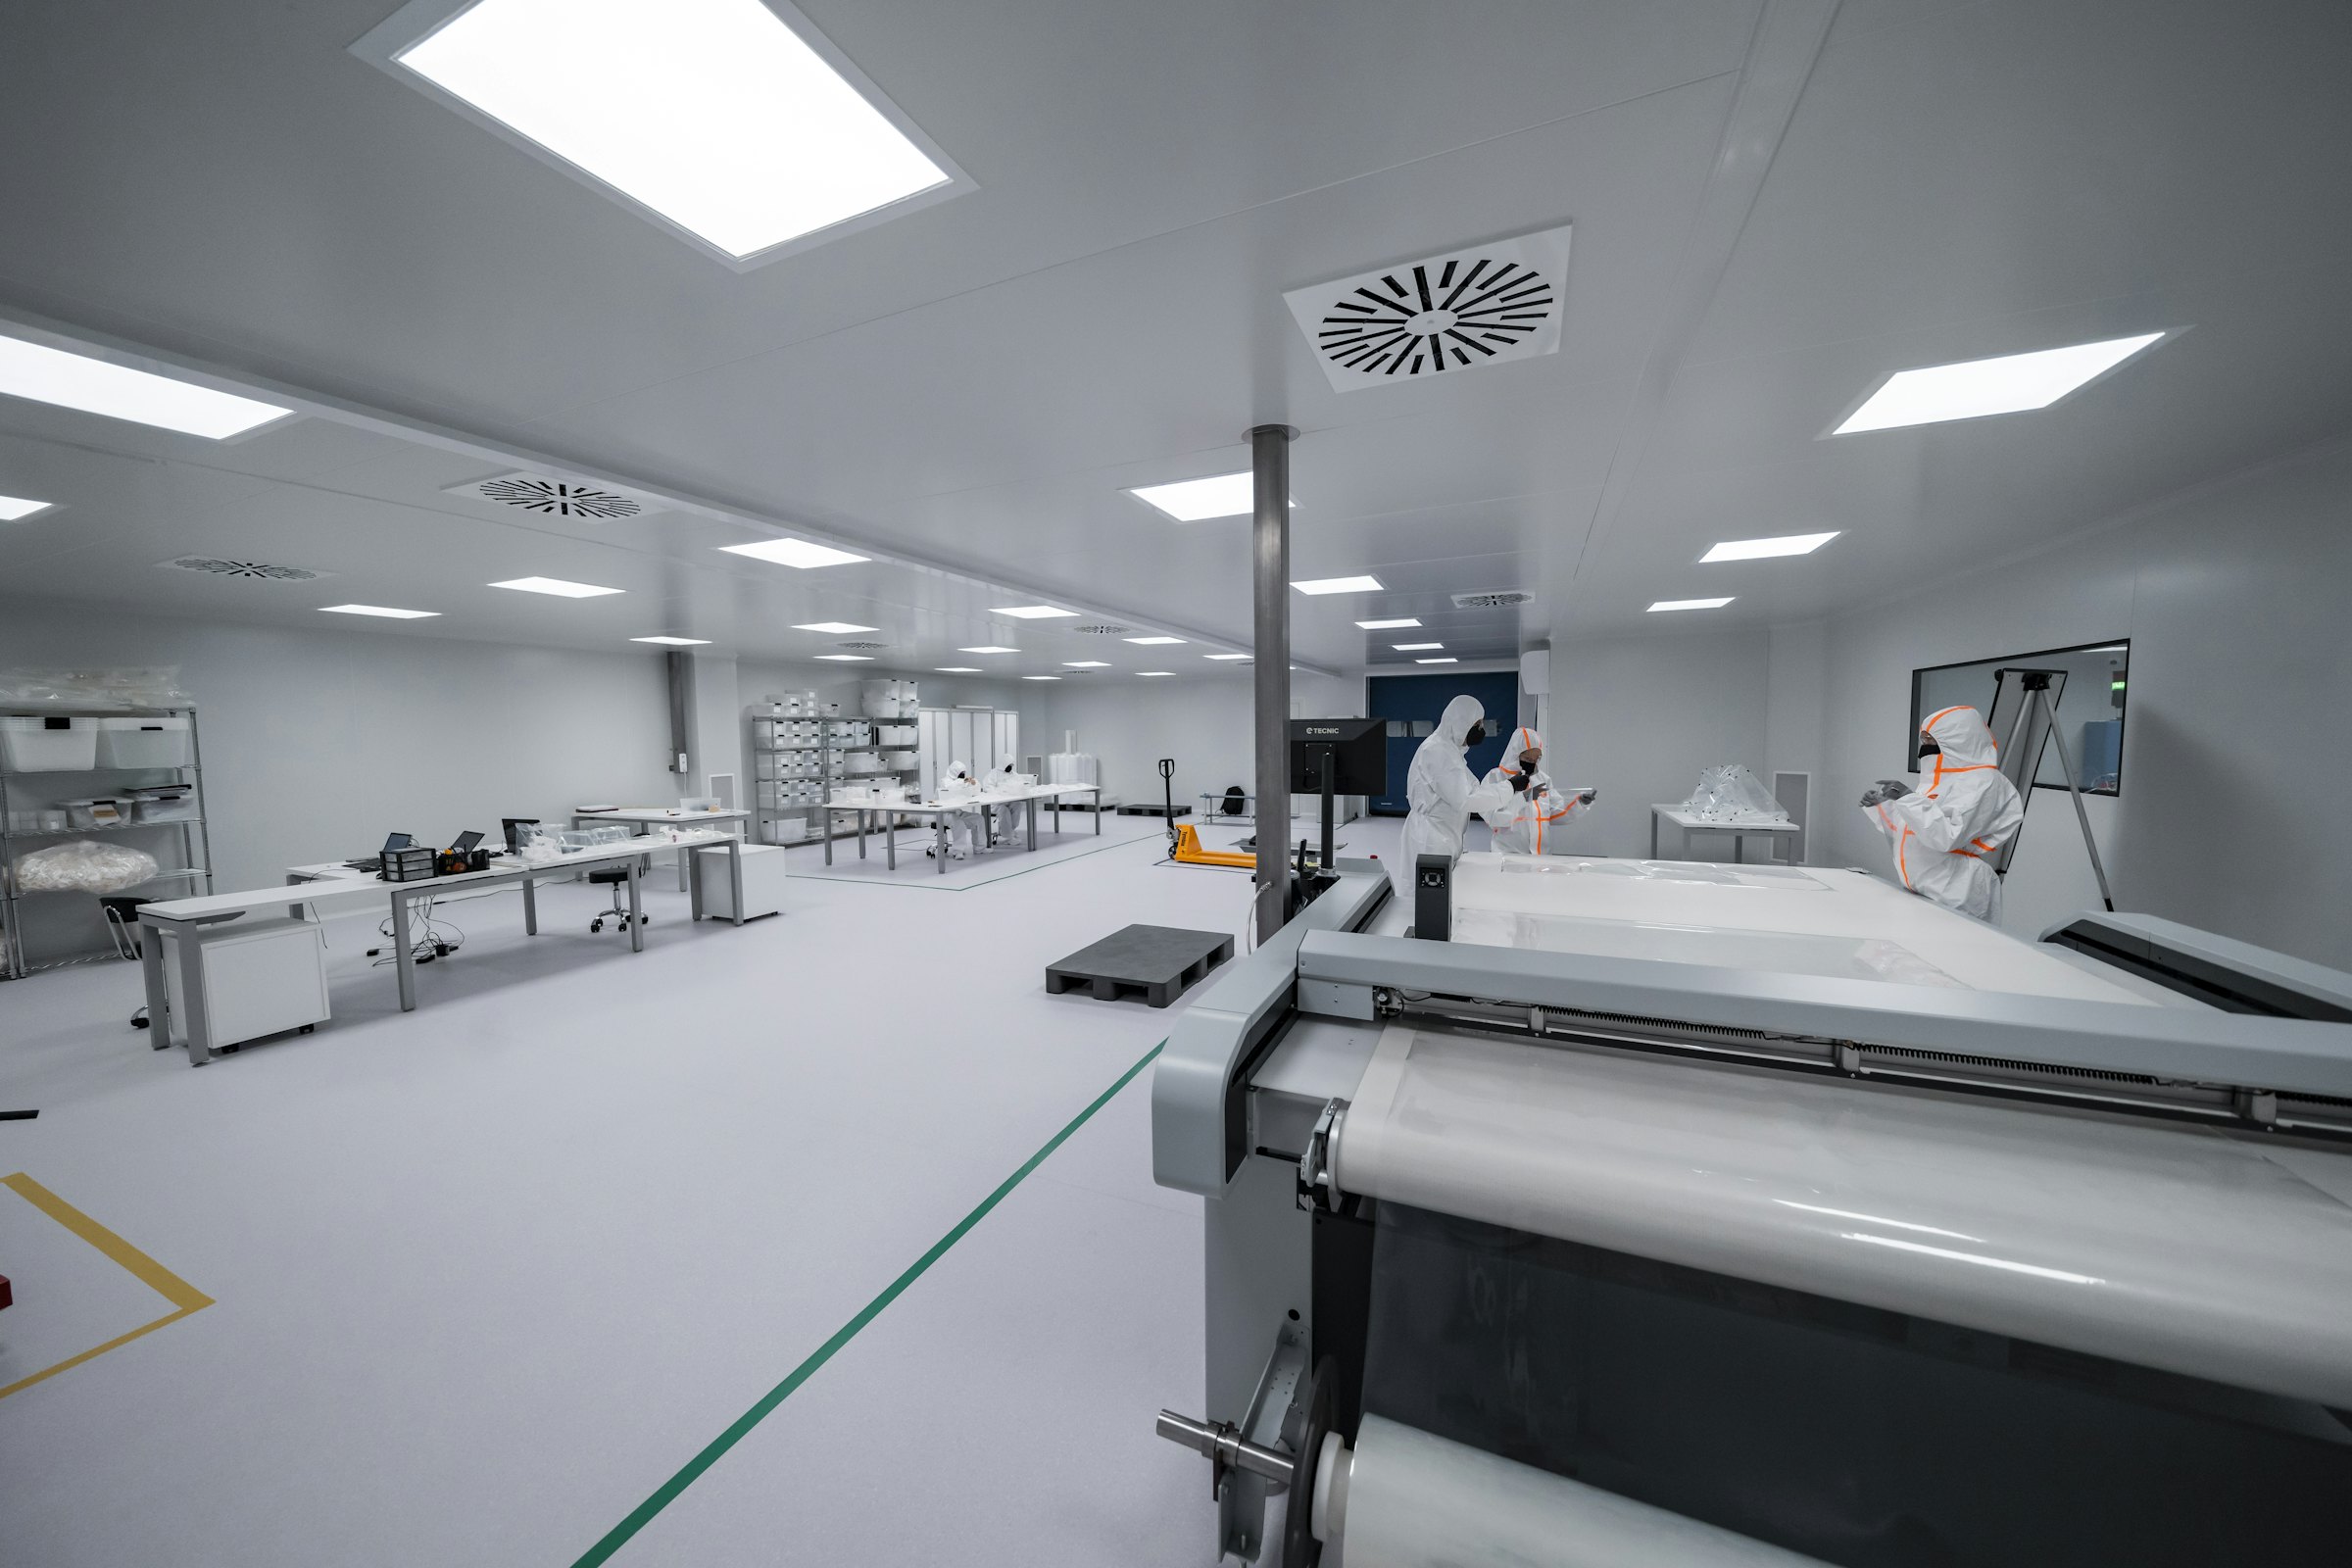
The distance between the binder and the floor
Most quality failures are not failures of intent. The procedure exists, the policy is approved, the training record is signed, and the practice on the floor still does not match what the document says. That distance is what an investigator finds, and it is what a warning letter gets written about.
Mrovin closes that distance. We work as practitioners rather than advisors: we hold the pen on the protocol, run the investigation, and sit in the room during the inspection. Every engagement ends in an executed, approved record that survives review.
- 15+ years
- in regulated manufacturing
- 10+ frameworks
- in active regulatory practice
- 500+ documents
- QMS integrated across acquisitions
- 50 systems
- qualified through IQ / OQ / PQ
Three ways we work with manufacturers
Every engagement is scoped to a defined deliverable, a named accountable owner, and a record that survives inspection. We do not hand over advice; we hand over executed, approved, defensible documentation.
-
Consulting
Gap assessments, remediation strategy and hands-on execution, led by practitioners who have held the pen on the documents an investigator asks to see.
- Regulatory gap assessment
- Remediation strategy & sequencing
- QMS build-out and integration
- Interim quality leadership
-
Validation & Qualification
Protocols authored, executed, deviated and reported to a defensible, ALCOA+ compliant standard, written to be run by your people.
- Facility, utility & equipment qualification
- Process validation and PPQ
- Cleaning and test method validation
- CSV and Part 11 data integrity
-
Training & Inspection Readiness
We prepare the people who will be in the room, not just the binders on the shelf.
- Role-based GxP and SOP training
- Mock FDA / notified body inspections
- 483 and observation response
- CAPA investigation coaching
The control areas regulators actually examine
Seven areas account for nearly every observation we are called in to answer. This is what we do inside each one.
-
Quality Management Systems
The system of record every other control depends on: document control, training, records, internal audit and management review. We build a QMS from nothing, remediate one that has drifted from practice, and integrate separate systems after an acquisition without losing traceability on either side.
-
Regulatory Affairs
Determining the pathway before money goes into the wrong one. We establish classification and route to market, assemble the submission or technical file, and manage the questions that come back: 510(k), De Novo, EU MDR technical documentation and international registrations.
-
Product Realization & Design Controls
A design history file an investigator can follow from user need to released product. Design inputs and outputs, verification and validation protocols, design reviews, transfer to manufacturing, and a traceability matrix that actually closes.
-
Supplier Controls
Your supplier’s nonconformance becomes your recall. We qualify and audit suppliers, write purchasing controls and quality agreements that bind change notification, and set incoming inspection to match the risk a component actually carries.
-
Manufacturing, Production & Process Controls
Proving the process does what you say it does, every time. Facility, utility and equipment qualification through IQ / OQ / PQ, process validation and PPQ, cleaning validation, test method validation, and the in-process controls that hold the validated state between runs.
-
Quality System Controls
The feedback loop regulators examine first. Nonconformance and CAPA investigations that reach real root cause, complaint handling, MDR and vigilance reporting inside the clock, and change control that keeps the validated state intact.
-
Risk Management
Risk as a live file, not a document written once to satisfy an auditor. Hazard analysis and FMEA to ISO 14971, benefit-risk determination, and post-market surveillance fed back into the file so the risk profile reflects the product as it behaves in the field.
What we work to
- 21 CFR Part 11 / 210 / 211
- 21 CFR Part 820 / 803 / 806
- ISO 13485
- ISO 14971
- EU MDR 2017/745
- EU GMP Annex 1 & Annex 11
- GAMP 5
- ALCOA+ data integrity
Tell us what you’re preparing for
The product, the target market, and where you are in the process. We come back with the pathway and the gaps that stand in it.
hello@mrovin.com